
 Do you have any questions? (088) 2080369 - 2345622 Pharmacy_QAAU@pharm.aun.edu.eg
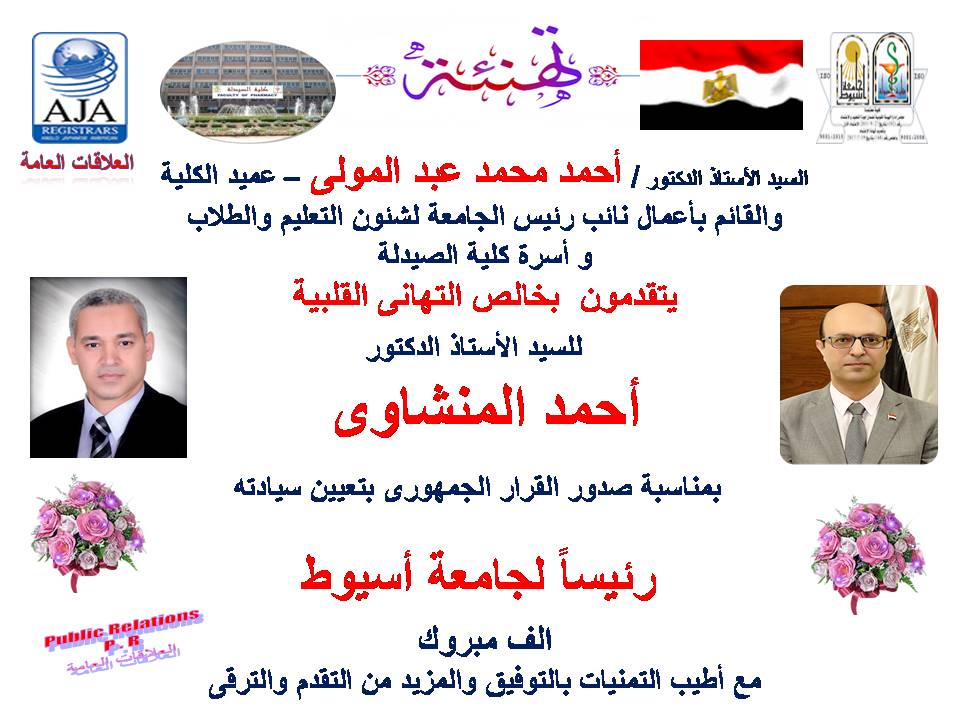
Prof. Dr. Ahmed Mohamed Abdel Mawla Dean of the faculty of Pharmacy and Acting Vice President for Education and Student Affairs and faculty family They extend their heartfelt congratulations For Prof. Dr Ahmed Al-Minshawi President of Assiut University
Impact of quercetin spanlastics on livin and caspase-9 expression in the treatment of psoriasis vulgaris
Research Abstract
Psoriasis is a chronic immune-mediated disease presented by erythematic lesions. It is an incurable disease; however, topical treatment attempts to stop the uncontrolled proliferation of skin cells. Quercetin (Qc) is a natural compound plentiful in several natural plant species and plays a vital role in treating psoriasis; however, it has poor aqueous solubility and penetrability. Consequently, our work aims to enhance Qc's dissolution and anti-psoriatic activity. In this work, Qc was isolated from Psidium guajava leaves, and spanlastics were prepared using the ethanol injection method. The % encapsulation efficiency (%EE) of Qc ranged from 76.40 ± 1.42 to 98.29 ± 0.2,4, and the Particle size (P.size) varied between 414.66 ± 1.94 to 748.33 ± 5.19 nm. In vitro release studies of Qc from spanlastic preparations were significantly higher than those from free drug dispersion. Selected Qc spanlastics formulation was included in Sodium carboxymethylcellulose (SCMC) 3.5% gel for biological application. Clinical improvement after eight weeks of treatment was powerfully distinguished. By the end of 8 weeks of treatment, Psoriasis Area and Severity Index (PASI) score decreased from 5.19 ± 1.14 at baseline to 2.26 ± 1.1. Qc is famous for its apoptotic-inducing activity, livin is one of the Inhibitors of apoptosis proteins (IAPs), and its function is mediated through the inhibition of caspase activation. Quantitative real-time polymer chain reaction (qrt- PCR) declared that the expression of livin (p = 0.002) was lowered while that of caspase-9 (P < 0.001) was elevated in comparison to those before treatment with QC spanlastics; thus, Qc spanlastics are promising easily prepared formulation for the treatment of psoriasis.
Research Date
Research Department
Research Journal
Journal of Drug Delivery Science and Technology
Research Member
Research Publisher
Elsevier
Research Rank
international
Research Vol
76
Research Year
2022
Meeting of the committee for community and environmental development at the Faculty of Pharmacy on Monday, February 13, 2023, at 10:00 AM

God willing, A meeting of the committee for community and environmental development will hold on Monday, February 13, 2023, at 10:00 AM
In the office of Vice Dean for Community Services and Environmental Development Affairs.
news category
خبر عام
A meeting of the laboratories and scientific equipment committee at the Faculty of Pharmacy on Monday, February 13, 2023 D at eleven (am)

God willing, the laboratories and scientific equipment committee will hold its meeting on Monday, February 13, 2023 at eleven (am)
in the office of Vice Dean for Community Services and Environmental Development Affairs.
news category
خبر عام
Design, Synthesis, Molecular Modeling, and Anticancer Evaluation of New VEGFR-2 Inhibitors Based on the Indolin-2-One Scaffold
Research Abstract
A new series of indoline-2-one derivatives was designed and synthesized based on the essential pharmacophoric features of VEGFR-2 inhibitors. Anti-proliferative activities were assessed for all derivatives against breast (MCF-7) and liver (HepG2) cancer cell lines, using sunitinib as a reference agent. The most potent anti-proliferative derivatives were evaluated for their VEGFR-2 inhibition activity. The effects of the most potent inhibitor, 17a, on cell cycle, apoptosis, and expression of apoptotic markers (caspase-3&-9, BAX, and Bcl-2) were studied. Molecular modeling studies, such as docking simulations, physicochemical properties prediction, and pharmacokinetic profiling were performed. The results revealed that derivatives 5b, 10e, 10g, 15a, and 17a exhibited potent anticancer activities with IC50 values from 0.74–4.62 µM against MCF-7 cell line (sunitinib IC50 = 4.77 µM) and from 1.13–8.81 µM against HepG2 cell line (sunitinib IC50 = 2.23 µM). Furthermore, these compounds displayed potent VEGFR-2 inhibitory activities with IC50 values of 0.160, 0.358, 0.087, 0.180, and 0.078 µM, respectively (sunitinib IC50 = 0.139 µM). Cell cycle analysis demonstrated the ability of 17a to induce a cell cycle arrest of the HepG2 cells at the S phase and increase the total apoptosis by 3.5-fold. Moreover, 17a upregulated the expression levels of apoptotic markers caspase-3 and -9 by 6.9-fold and 3.7-fold, respectively. In addition, 17a increased the expression level of BAX by 2.7-fold while decreasing the expression level of Bcl-2 by 1.9-fold. The molecular docking simulations displayed enhanced binding interactions and similar placement as sunitinib inside the active pocket of VEGFR-2. The molecular modeling calculations showed that all the test compounds were in accordance with Lipinski and Veber rules for oral bioavailability and had promising drug-likeness behavior.
Research Date
Research Journal
Pharmaceuticals
Research Member
Research Publisher
MDPI
Research Vol
15
Research Website
https://doi.org/10.3390/ph15111416
Research Year
2022
New 1,2,3-triazole linked ciprofloxacin-chalcones induce DNA damage by inhibiting human topoisomerase I& II and tubulin polymerization
Research Abstract
A series of novel 1,2,3-triazole-linked ciprofloxacin-chalcones 4a-j were synthesised as potential anticancer agents. Hybrids 4a-j exhibited remarkable anti-proliferative activity against colon cancer cells. Compounds 4a-j displayed IC50s ranged from 2.53-8.67 µM, 8.67–62.47 µM, and 4.19–24.37 µM for HCT116, HT29, and Caco-2 cells; respectively, whereas the doxorubicin, showed IC50 values of 1.22, 0.88, and 4.15 µM. Compounds 4a, 4b, 4e, 4i, and 4j were the most potent against HCT116 with IC50 values of 3.57, 4.81, 4.32, 4.87, and 2.53 µM, respectively, compared to doxorubicin (IC50 = 1.22 µM). Also, hybrids 4a, 4b, 4e, 4i, and 4j exhibited remarkable inhibitory activities against topoisomerase I, II, and tubulin polymerisation. They increased the protein expression level of γH2AX, indicating DNA damage, and arrested HCT116 in G2/M phase, possibly through the ATR/CHK1/Cdc25C pathway. Thus, the novel ciprofloxacin hybrids could be exploited as potential leads for further investigation as novel anticancer medicines to fight colorectal carcinoma.
Research Date
Research Department
Research Journal
Journal of Enzyme Inhibition and Medicinal Chemistry
Research Member
Research Publisher
Taylor & Francis Online
Research Vol
37
Research Website
10.1080/14756366.2022.2072308
Research Year
2022
Nicotinonitrile as an essential scaffold in medicinal chemistry: an updated review (2015– present)
Research Abstract
Many studies have been carried out to find synthetic pathways to nicotinonitriles (3-cyanopyridines) and their analogs, which has resulted in a substantial amount of knowledge on their biological, therapeutic, and medical characteristics. Some of nicotinonitrile derivatives have emerged as marketed drugs such as bosutinib, milrinone, neratinib, and olprinone. In this review, we discuss the recently synthesized nicotinonitrile-based hybrids that possess important biological activity and will be useful for drug discovery and development.
Research Date
Research Department
Research Journal
Journal of advanced Biomedical and Pharmaceutical Sciences
Research Member
Research Publisher
JABPS
Research Vol
6
Research Website
10.21608/jabps.2022.162396.1166
Research Year
2023